Regulatory and QMS lead for medical software teams
I work inside your product team to qualify and classify your software, build an ISO 13485 quality system that fits how you ship, and prepare the technical documentation your Notified Body will assess — for the EU, the UK and Southeast Asia.
- Published prices
- Independent consultant, not a Notified Body
- Reply within 1 business day
- Independent of assessorsI prepare you for assessment. Certificates are issued by Notified Bodies, never by a consultant.Who does what →
- Prices on the siteStrategy call from €200 · Gap analysis about €3,000 · ISO 13485 QMS from €5,000 · EU MDR support €10,000–25,000.Compare services →
- Named recommendationsEight LinkedIn recommendations from people who worked with me, with links to their profiles.Read them →
- Guides with sourcesPractical guides on EU MDR, ISO 13485, the UK and ASEAN, each listing the official documents behind it.Browse guides →
Where are you now?
Pick the closest match. Each one leads to the service that fits it, with scope, timing and price.
“Is our software a medical device, and which class?”
Qualification under MDR Article 2(1) and MDCG 2019-11, classification under Rule 11, and a route to market you can plan and budget.
Regulatory strategy & classification“We need a QMS that doesn’t slow down our sprints.”
An ISO 13485 quality system built around your Jira or GitHub workflow, with IEC 62304 and ISO 14971 built in.
ISO 13485 QMS implementation“Our technical file is drafted. Is it ready for the Notified Body?”
An independent review against MDR Annexes I, II and III with findings rated by severity and an action plan.
Technical documentation gap analysis“We are preparing for, or already in, Notified Body review.”
Hands-on support to complete the technical documentation, prepare for audits and answer the Notified Body’s questions.
EU MDR consulting“We have CE marking and want Singapore or Malaysia.”
Plan HSA and MDA submissions that reuse your EU MDR evidence, and the Malaysia verification route for HSA-registered devices.
Singapore & Malaysia registration“We are on the market and have no QA/RA lead.”
Part-time support for change assessments, post-market surveillance, CAPA, internal audits and surveillance audits.
Ongoing QA/RA supportPrefer to build the QMS yourselves? See the ISO 13485 template set for software companies.
What you can buy
Every engagement starts with a free call. Packages have set prices; technical writing and ongoing support are charged by the hour. Timings below are for my part of the work; assessment by a Notified Body or regulator comes on top.
EU MDR consulting for medical software
Classification, technical documentation to Annexes II and III, clinical evaluation, Notified Body preparation and audit support.
€10,000–25,000 fixed feeTypically 6–9 months to Notified Body submission (Class IIa), depending on your starting point
Scope and stages → Quality systemISO 13485 QMS for software teams
Procedures, records and training for a QMS that works with sprints, pull requests and CI/CD.
€20,000 tailored · €30,000 from scratch3 or 6 months; template package €5,000, 1 month
Scope and stages → ReviewTechnical documentation gap analysis
Independent review of your existing technical file before submission, with severity-rated findings and an action plan.
About €3,0001–4 weeks, depending on scope
What you get → Southeast AsiaSingapore & Malaysia registration
Route planning and dossier preparation for HSA and MDA, built on the documentation you already have for the EU.
Quote after scopingDepends on device class, route and the state of your EU documentation
How it works → Time and materialsOngoing QA/RA support
A part-time regulatory and quality lead at any stage, from development and V&V to post-market, without a full-time hire.
€50/hour, time and materialsDesign and development, V&V, maintenance and change management
What’s included → Do it yourselfISO 13485 QMS templates
50+ editable documents for a software company’s QMS, with IEC 62304, ISO 14971 and IEC 62366-1 integrated.
€5,000Template package, 1 month
Contents →Regulations and standards I work with
EU MDR 2017/745ISO 13485ISO 14971IEC 62304IEC 62366-1MDCG 2019-16 cybersecurityEU AI ActUK MDR 2002HSA SingaporeMDA Malaysia
Who does what
Under EU MDR the manufacturer is responsible for the device, and the Notified Body decides whether to issue a certificate. My job is to make your team ready for that decision. No consultant can guarantee a certificate or a certificate date.
| Activity | Your team (manufacturer) | Me (consultant) | Notified Body |
|---|---|---|---|
| Intended purpose, qualification and class | Decides and signs off | Prepares rationale and options | Reviews during assessment |
| QMS procedures and records | Approves, uses, keeps records | Drafts and adapts, trains, audits internally | Audits the QMS |
| Design, code, testing and test evidence | Does the work | Defines required evidence, reviews | Samples and assesses |
| Technical documentation (Annexes II and III) | Provides content, approves | Structures, writes and reviews sections | Assesses |
| Clinical evaluation | Provides clinical input and data, approves | Plans and drafts with your clinical experts | Assesses |
| Certificate decision | — | — | Decides |
| Placing on the market, vigilance, PRRC | Responsible | Sets up processes, supports | Surveillance audits |
Exact split is written into each proposal. For Class I software without a measuring function, there is usually no Notified Body: the manufacturer declares conformity itself.
From first call to a written proposal
Free call (30 minutes)
You describe the product, intended use, markets and deadlines. I tell you what I see as the likely route and the main risks, whether or not we work together.
Scoping
If needed, I look at your existing documents and agree with you what is in scope, what your team will do and what the Notified Body or regulator will do.
Written proposal
The price for the agreed scope, the deliverables, the stages and what I need from you at each stage.
Work inside your team
I join your planning and reviews, turn requirements into tasks your developers can implement, and prepare you for each audit or submission.
Projects I have worked on
These projects were delivered in roles I held before founding TrustedTraceMed. Each case states what I did and what the client team did. Client names are not shown.
Haemodialysis patient management SaMD
ISO 13485 QMS built from scratch for the manufacturer, and technical documentation aligned with the development partner.
Read the case → AI/ML SaMD · UK and EUAI-assisted dental X-ray software
QMS implementation for the legal manufacturer and review of deliverables from three suppliers for one technical file.
Read the case → eHealth · GDPR · CroatiaNational eHealth platform
Compliance oversight for EMR, ePrescription, pharmacy and laboratory modules from discovery to production.
Read the case →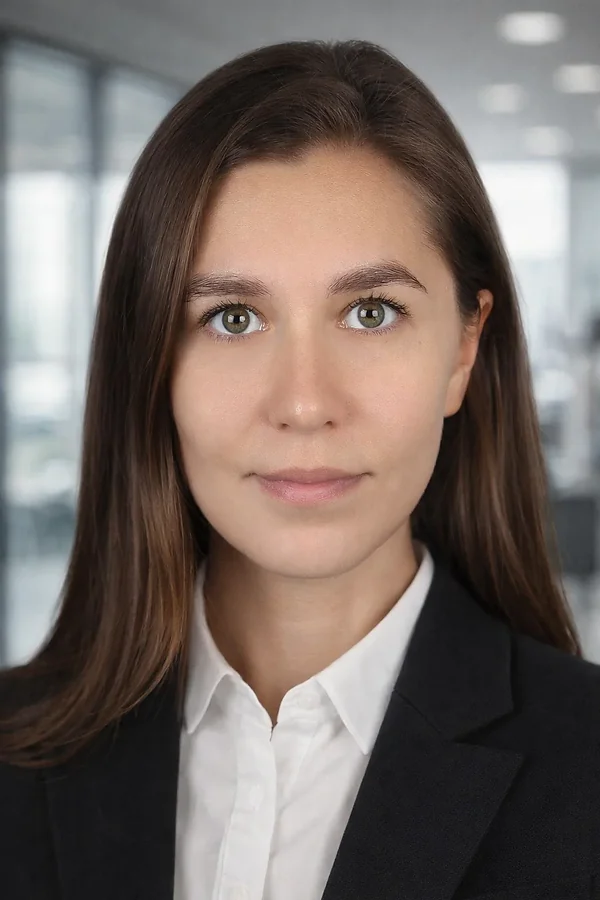
Lizaveta Dabrynskaya
I have spent more than 13 years in the healthcare domain, on X-ray systems, clinical trial platforms, national eHealth platforms and AI-based diagnostic software. In my last role before founding TrustedTraceMed, I built an ISO 13485 QMS from scratch for a Class IIa SaMD, wrote the SOPs, ran risk management (ISO 14971) and usability engineering (IEC 62366-1), and was the main contact for the Notified Body and audit companies.
I also come from product management and business analysis, so I write requirements and user stories as comfortably as SOPs. Your project is run by me personally — there is no hand-over to junior staff.
What people who worked with me say
Recommendations published on LinkedIn. Each name links to the author’s profile; job titles are as shown there.
She takes her job seriously and always approaches problems in a systematic way. Always there to clarify requirements and offer assistance no matter the hour.
 Patrik Glavica
Security Master & Team Lead · Ericsson Nikola TeslaLinkedIn recommendation
Her work ethics are of the highest order — always on top of tasks, very proactive, well organised and willing to help. She has my highest recommendation.
Director, Compliance Assurance · EPAM SystemsLinkedIn recommendation
Lizaveta is very knowledgeable in her field and open to learning new things. She is organised and won't let go until project results are achieved with success.
 Sherif S.
Business Development ManagerLinkedIn recommendation
In 2–3 weeks she started defining full modules by herself, exceeding personal goals every time. Competent, calm, organised — excellent potential and a great team player.
 Marko Kranjčić
Technical Coordinator · Ericsson Nikola TeslaLinkedIn recommendation
I cannot emphasise enough how big Liza's contribution was on our national healthcare information system. Top professional, remarkable attention to detail. Cannot recommend her enough.
Solution Manager · Ericsson Nikola TeslaLinkedIn recommendation
Together we developed SaMD and obtained Malaysian GDPMD/MDA certificates. Lisa guided the entire team, assisted with usability risk management and documentation. Highly skilled and a pleasure to work with.
 Alex Nosach
Senior UX/UI DesignerLinkedIn recommendation
Lizaveta explained what we were working on across multiple SaMD certifications. She conducted training sessions, presented demos to the client and managed documentation. The client feedback was excellent.
 Stanislav Khytrych
Software Architect & Lead Frontend DevLinkedIn recommendation
Only thanks to Liza's guidance did we successfully obtain ISO 13485/9001 and MDA certifications for the Malaysian market. Her attention to detail and proactive approach ensured all compliance goals were met.
Lead BA/SA in Healthcare, Product ManagerLinkedIn recommendation
Read before you decide
Free, practical guides. Each one lists the official sources it relies on.
Is your software a medical device under EU MDR?
Qualification under Article 2(1) and MDCG 2019-11, and where classification under Rule 11 comes in.
Read the guide →How long EU MDR certification takes for software
Your preparation time and the Notified Body’s assessment time, planned separately and realistically.
Read the guide →EU MDR technical documentation for SaMD
What Annexes II and III expect from a software device, section by section.
Read the guide →Frequently asked questions
No. I am an independent consultant. Under EU MDR, certificates are issued by Notified Bodies; UK approved bodies assess devices for UKCA marking; in Singapore and Malaysia, HSA and MDA register devices. I help you prepare: classification, quality system, technical documentation and audit readiness. The manufacturer remains responsible for the device.
My published prices: strategy call €200 (1.5-hour workshop) or €1,000 with analysis of your input; gap analysis about €3,000; ISO 13485 QMS implementation €5,000 (template package), €20,000 (tailored, 3 months) or €30,000 (built from scratch, 6 months); EU MDR consulting €10,000–25,000; technical writing €35 per hour and ongoing support €50 per hour. Fees are in EUR, excluding taxes. Notified Body fees, testing and other third-party costs are separate. All prices.
Plan two separate phases. Preparation: for a Class IIa SaMD, my consulting workstream to a submission-ready technical file typically takes 6–9 months, depending on your starting point. Assessment: in the European Commission’s 2026 survey of Notified Bodies, 51% reported 13–18 months from signed written agreement to certificate for QMS and product certification, and a further 41% reported longer. Nobody can promise you a certificate date. How to plan the timeline.
Yes. I have worked on AI-based diagnostic software, including image analysis. Typical topics are intended purpose and classification, data and algorithm validation evidence, clinical evaluation, change control for model updates, and how the EU AI Act will apply to your device.
I reply by email within one business day to agree a time for a free 30-minute call. There is no automatic booking: we pick a time together by email. You can tell me in the form what you would like to cover, so the call starts from your questions.
Tell me where your product is today
A free 30-minute call to discuss your route, risks and next steps. No commitment.